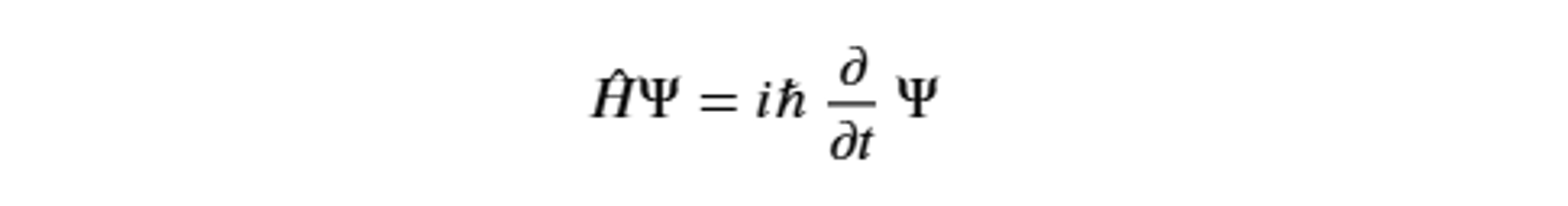
Home Authors Posts by staff

staff

112 POSTS 0 COMMENTS

Gene and cell therapies, market trends for Q4 2025

0

The latest figures on the gene and cell therapy market released by the Alliance for Regenerative Medicine for the fourth quarter of 2025 show a clear lead for the United States over Europe, both in the number of clinical trials (916 vs 337) and in total investments ($2.5 billion vs $0.6 billion), despite a nearly identical number of developers (703 vs 685).

On the regulatory front, the new “plausible mechanism pathway” announced by the FDA at the end of last year is designed to accelerate access to personalized gene editing therapies and other treatments targeting unmet medical needs. Also in the United States, Baby KJ became the first child to receive an in vivo gene editing treatment for the ultra-rare CPS1 deficiency—a condition caused by the lack of the carbamoyl phosphate synthetase 1 enzyme, leading to toxic ammonia accumulation in the body. The CRISPR-based therapy was developed in just six months from diagnosis, with FDA authorization granted in only one week.

In Europe, meanwhile, the publication of the Biotech Act by the European Commission on December 16, 2025, is being seen as a turning point for the biotech medicines sector. The initiative combines financial incentives with regulatory improvements aimed at supporting the development of gene and cell therapies.

European pipelines lag behind

The report also highlights a robust U.S. pipeline, with at least twenty gene and cell therapy products already approved (three in 2026) or nearing regulatory submission.

By contrast, Europe’s clinical pipeline remains significantly smaller, with only five products approaching or having reached approval in 2026. Among them is the gene therapy developed by Fondazione Telethon for Wiskott-Aldrich syndrome, approved by the European Commission on January 16, 2026. The therapy had also received FDA approval in December 2025 and marks a milestone as the first case worldwide of a gene therapy for a rare genetic disease fully developed by a non-profit organization. Two additional approvals are expected in Europe this year, involving a cell therapy for metastatic melanoma and a gene therapy for retinitis pigmentosa.

On the corporate front, the quarter was marked by Bristol Myers Squibb’s $1.5 billion acquisition of Orbital Therapeutics, alongside several new strategic partnerships. Notably, Italy’s Chiesi has entered into a collaboration with Arbor Biotechnologies focused on genome editing for the treatment of primary hyperoxaluria type 1, with an upfront commitment of $115 million from Chiesi.
EMA strengthens stakeholder dialogue across the medicines lifecycle

0

The latest report on EMA’s stakeholder engagement activities for 2024–2025 highlights an intensification of dialogue and collaboration aimed at integrating external contributions into both the evaluation and safety monitoring of medicines. The Agency confirms a structured and increasingly inclusive approach, designed to ensure that multiple perspectives are reflected throughout regulatory processes.

At the core of this strategy is a multi-stakeholder model that continues to evolve into a dynamic platform for exchange. Within this framework, patient-centricity has consolidated its role as a guiding principle, consistently considered across all phases of a medicinal product’s lifecycle—from development to post-market surveillance.

Growing participation and structured collaboration

One of the most tangible signals of this strengthened engagement is the increase in participation within EMA’s key working groups. Representatives from patient and consumer associations involved in the Patients’ and Consumers’ Working Party rose from 22 to 25 member organisations, reflecting a broader inclusion of patient voices. A similar upward trend was observed in the Healthcare Professionals’ Working Party (HCPWP), confirming the Agency’s effort to expand the contribution of healthcare professionals in regulatory discussions.

Engagement also extended to the academic and research community. Independent and not-for-profit research was supported through dedicated dialogue channels such as Academia Briefing Meetings (ABMs) and Innovation Task Force (ITF) meetings. These initiatives were complemented by concrete support measures, including fee reductions for scientific advice, the qualification of novel methodologies, and procedural assistance—tools aimed at fostering innovation while maintaining regulatory rigor.

Industry engagement and regulatory alignment

Parallel to this, EMA maintained regular exchanges with industrial stakeholders. Interactions took place both within the Industry Standing Group (ISG) and through bilateral meetings with individual organisations. These discussions focused primarily on the implementation of new legislative requirements, as well as on assessing the broader impact of cross-sectoral regulations, an increasingly relevant aspect in a complex and interconnected regulatory landscape.

Overall, the report outlines a stakeholder engagement model that is becoming more structured, participatory and aligned with the evolving needs of the European healthcare ecosystem.

ECHA’s proposed PFAS restrictions

0

In March 2026, the Risk Assessment Committee (RAC) and the Socio-Economic Analysis Committee (SEAC) of the European Chemicals Agency (ECHA) launched a public consultation on their opinions regarding a potential EU-wide restriction on PFAS. The proposed measure would limit the manufacture, placing on the market, and use of these substances, while allowing for specific derogations. Stakeholders can submit comments until 25 May 2026.

Growing risks for human health and the environment

In its final opinion, RAC highlights that PFAS pose increasing risks to both human health and the environment due to their high persistence. According to the Committee, existing regulatory measures are not sufficient to effectively control emissions, making coordinated EU-level action necessary. Where derogations are granted, RAC recommends a set of accompanying measures, including:

  • site-specific PFAS management plans for manufacturers and industrial users;
  • clear consumer labelling;
  • guidance on safe use and disposal.

SEAC’s draft opinion stresses that PFAS are widely used across multiple industrial applications in Europe, reinforcing the need for a harmonised EU approach to prevent internal market distortions. The Committee suggests that derogations should be limited to uses where no viable alternatives are currently available, alongside risk management measures to minimise emissions.

EFPIA raises concerns over pharmaceutical impact

The European Federation of Pharmaceutical Industries and Associations (EFPIA) responded to the committees’ opinions by highlighting the absence of explicit derogations for the pharmaceutical sector. According to EFPIA, “the absence of clear and durable derogations for medicines would compromise patient access to critical treatments and drive manufacturing out of Europe.”

The organisation notes that PFAS materials are essential in several pharmaceutical manufacturing processes, including the production of active substances and delivery devices. Moreover, replacing PFAS components is not straightforward, as it would require extensive R&D efforts as well as legal and regulatory approvals.

IFPMA’s position on AI in pharmaceutical manufacturing

0

Artificial Intelligence (AI) and Machine Learning (ML) are playing an increasingly central role in pharmaceutical manufacturing and control. A position paper published by IFPMA in March 2026 outlines both the opportunities and the challenges associated with their adoption across the sector.

Enhancing efficiency and enabling new applications

According to the document, AI has the potential to accelerate decision-making processes and enhance operational efficiency, ultimately leading to more robust manufacturing processes and shorter cycle times. By automating repetitive tasks with higher accuracy, AI can also deliver significant benefits in terms of data integrity and analytical capabilities.

Several real-world applications are already emerging, including predictive and control models such as digital twins of manufacturing processes. In parallel, generative AI is being used for text-related activities, for example in the preparation of regulatory documentation.

Regulatory alignment and the need for a common framework

From a regulatory perspective, a risk-based approach to the implementation and updating of AI and ML systems has been adopted by many international authorities. The paper highlights a strong alignment between the requirements set by the European Medicines Agency (EMA) and the US Food and Drug Administration (FDA), suggesting a growing convergence at the global level.

IFPMA also points to the opportunity to develop a sector-specific AI risk framework for the pharmaceutical industry, alongside the promotion of global harmonisation of regulatory standards to support wider adoption in manufacturing.

Addressing risks and ensuring sustainable adoption

The document highlights several AI-specific risks that require careful management, including automation bias, the potential erosion of human oversight, challenges related to explainability and interpretability, autonomous learning, data quality, and lifecycle maintenance.

To address these challenges, the position paper proposes a set of actions aimed at enabling safe, effective and scalable implementation of AI and ML technologies across the pharmaceutical sector.

The role of quantum mechanics in drug discovery

0
Edited by Matteo Iafrate - Pharmaceutical Analytical Chemist

Quantum mechanics allows us to obtain very accurate results in the study of molecules. Over the years, numerous computational methods have been developed that are capable of studying the system under examination with different approaches, always rigorously based on quantum theories. Quantum mechanics attempts to interpret, with excellent results, through statistical probability, the behavior of matter and its energy at the atomic and subatomic level, differentiating itself from classical mechanics, which applies to macroscopic systems. It is important to underline that classical mechanics fails at the molecular level because several properties such as electron delocalization and chemical bonding are strictly linked to quantum effects. Furthermore, quantum mechanical methods offer high accuracy for small systems, while classical methods can handle large systems with reduced accuracy.

Theoretical Foundations

Some of the main theoretical aspects of quantum theory concern wave-particle duality, the quantization of energy, wave function and probability, the superposition principle, the Heisenberg uncertainty principle, and entanglement. The study of time evolution, which describes how the wave function changes over time, with a probabilistic approach, is described by the Schrödinger equation:

Where Ĥ represents the Hamiltonian operator (including kinetic and potential energy terms), and ψ the wave function

In scientific literature, the use of computational methods that make use of quantum mechanics is increasingly present. Among the most widely used are: density functional theory (DFT), Hartree-Fock (HF), quantum mechanics/molecular mechanics (QM/MM), and fragment molecular orbitals (FMO).

Methods and applications

These techniques are based on quantum mechanics, but operate and describe the system under study differently. For example, density functional theory (DFT) is based on the electron density, while the Hartree-Fock method (HF) is based on the wave function. Quantum mechanics/molecular mechanics (QM/MM) is a hybrid method, aiming to combine the accuracy of quantum mechanics  for the molecular regions under study with the efficiency of molecular mechanics (MM). The fragmented molecular orbital method (FMO) splits a large molecule into smaller fragments, with the aim of treating each fragment quantum mechanically and taking into account their interactions.

Many of the results obtained, with the integration of quantum computing, aim to accelerate the drug development cycle and improve workflows, model interactions between small molecules and macromolecules, understand the functioning of enzymes based on electronic effects, model protein-ligand interactions and much more. The increase in computational performance and the excellent results obtained identify the quantum mechanical approach as very promising in drug discovery. In the coming years, computational drug discovery combined with quantum mechanics is expected to become increasingly common.

References

  • Sarfaraz K. Niazi. Quantum Mechanics in Drug Discovery: A Comprehensive Review of Methods, Applications, and Future Directions, Int. J. Mol. Sci., 2025.
  • Yidong Zhou, Jintai Chen, Jinglei Cheng, Xu Cao, Yuanyuan Zhang, Gopal Karemore, Marinka Zitnik, Frederic T. Chong, Junyu Liu, Tianfan Fu, Zhiding Liang. Quantum-machine-assisted drug discovery, npj Drug Discovery, 2026.

Be Ready, the new partnership for pandemic preparedness

0

The new partnership, Be Ready, launched by the European Commission in February 2026, is designed to significantly strengthen and expand research activities dedicated to pandemic preparedness across Europe. Under the initiative, a total budget of €120 million from the Horizon Europe programme will be made available to support projects focusing on emerging and re-emerging pathogens, with the goal of accelerating the development of advanced diagnostics, innovative treatments and next-generation vaccines. The partnership is coordinated by the French research authority ANRS-MIE and brings together a broad network of 81 organisations from 27 countries, reflecting a strong commitment to international collaboration.

Beyond funding, Be Ready will work to align national research priorities among participating countries, fostering greater strategic coherence and reducing fragmentation. It will also enhance coordination in pandemic preparedness research through the definition of a shared strategic and research innovation agenda, enabling a more structured and forward-looking approach.

Joint transnational calls will play a key role in supporting multinational research projects, promoting coordinated funding mechanisms among national and European funders. At the same time, all core components of the initiative will be fully integrated into the wider research ecosystem, ensuring that Be Ready remains active and effective not only during health emergencies but also in the periods between epidemics, strengthening long-term resilience.

Reactions to the EU Court of Justice’s rejection of the annulment of the UWWTD

The European General Court has dismissed EFPIA’s legal action (Case T-158/25) seeking the annulment of the Urban Wastewater Treatment Directive (UWWTD). The ruling, however, is based solely on procedural grounds and does not address the substance of the Directive, leaving the underlying legal questions unresolved.

Court dismisses EFPIA case on procedural grounds

In its decision, the Court concluded that EFPIA had not demonstrated that it was individually concerned by the provisions of the Directive it challenged. This requirement, together with being directly concerned, represents one of the cumulative legal conditions that must be met in order to establish legal standing before the EU General Court. Because these conditions were not considered satisfied, the Court declared the action inadmissible. As a result, the ruling does not examine the merits of EFPIA’s arguments nor the broader legal validity of the Urban Wastewater Treatment Directive.

Following the decision, EFPIA reaffirmed its commitment to ensuring that EU legislation is implemented in full respect of fundamental principles of EU law. Among these, the association highlighted the importance of the “polluter pays” principle, as well as the principles of non-discrimination and proportionality. Since the Court did not address the substance of the case, these legal issues remain open and could still be examined in other proceedings.

Implications for AESGP’s intervention request

The dismissal of EFPIA’s action also resulted in the non-adjudication of AESGP’s request to intervene in the case. AESGP, which represents the self-care industry, had sought to participate in the proceedings to support the arguments raised. According to the association, the key legal questions surrounding proportionality, non-discrimination and the polluter-pays principle should instead be assessed in other ongoing cases, particularly Poland’s legal challenge against the UWWTD (Case C-193/25).

Legal scrutiny of the Directive is continuing through other channels. In parallel with EFPIA’s action, the Irish Pharmaceutical Healthcare Association (IPHA) is advancing a case before the High Court in Ireland, seeking a referral to the Court of Justice of the European Union (CJEU) for a preliminary ruling on the validity of the Directive. The General Court’s order also clarifies an important procedural point: when national courts have doubts about the validity of an EU Directive, they should refer the matter to the CJEU.

For this reason, although EFPIA’s action has been dismissed, the broader legal debate surrounding the Urban Wastewater Treatment Directive is likely to continue in both national courts and EU-level proceedings.

Concept paper on the revision of Annex 15

0

As agreed during the 115th meeting of EMA’s GMP/GDP Inspectors Working Group (GMDP IWG) in September 2024, the European Medicines Agency has launched a targeted revision of Annex 15 of the Good Manufacturing Practices (GMPs). The consultation period on the Concept Paper, jointly published by EMA and PIC/S in February 2026, will remain open until 9 April 2026. This initial phase will be followed by the publication of a revised draft guideline and a second round of consultation, with the final text expected to be published by the end of 2026.

The initiative to revise Annex 15 stems from lessons learned since 2020 regarding the presence of N-nitrosamine impurities in medicines containing sartans as active ingredients. The report published on the case highlighted, among other aspects, an insufficient level of knowledge and understanding of the manufacturing processes of active substances by their producers. This lack of process understanding contributed to quality issues affecting active pharmaceutical ingredients (APIs) and to inadequate measures to control contamination by impurities. The report on the nitrosamine case ultimately recommended making the application of Annex 15 mandatory also for API manufacturers.

This recommendation was taken up in 2024 during the 115th meeting of the GMDP IWG and has now led to the publication of the new Concept Paper by EMA and PIC/S, the international scheme for cooperation in pharmaceutical inspections.

Objectives of the revision

According to the document, the revision aims to extend the scope of Annex 15 to manufacturers of active substances, both chemical and biological. The update will also introduce selected modifications to the current text and strengthen the link with the provisions set out in Part II of the EU GMP guidelines, which specifically address active substances. Another key objective is to align the Annex more closely with the ICH Q9 (R1) guideline on Quality Risk Management, in order to ensure greater consistency between the two frameworks when making decisions related to qualification and validation activities.

The Concept Paper indicates that greater emphasis should be placed on the validation documentation for active substances, including the Validation Master File, as well as on the definition of a clearer qualification and validation policy. Such a policy should specify roles and responsibilities, the criteria used, including acceptance criteria, and the monitoring strategies adopted during the lifecycle of the manufacturing process.

A lifecycle-based approach to validation

At the core of the proposed revision is the introduction of a lifecycle-based approach to validation. This approach would cover the entire process lifecycle, from process design and qualification to ongoing performance monitoring and final evaluation, based on data and metrics collected throughout the manufacturing lifecycle.

The Concept Paper also highlights the need for robust procedures to investigate non-conforming results, supported by appropriate corrective and preventive actions (CAPA). In the area of qualification, the revision is expected to strengthen several key concepts, including the definition of User Requirements Specifications (URS) and the role of Factory Acceptance Testing and Site Acceptance Testing (FAT/SAT).

EMA committees’ 2026 work plans: regulatory priorities and strategic focus

The European Medicines Agency (EMA) has outlined its 2026 priorities through the annual work plans of its scientific committees, marking a year that will be crucial for regulatory transition and innovation. A central element will be the preparation for the implementation of the new EU pharmaceutical legislation. The Committee for Medicinal Products for Human Use (CHMP) will focus on pre-authorisation activities, initial evaluations, specialised scientific areas and horizontal initiatives, ensuring readiness for the evolving legal framework.

Orphan medicinal products and pharmacovigilance

The Committee for Orphan Medicinal Products (COMP) will continue its evidence-based assessment of orphan designations and maintenance of orphan status. Planned activities also include work on indirect comparisons and the integration of new approach methodologies (NAMs), reinforcing scientific robustness in regulatory decision-making.

Pharmacovigilance will remain high on the agenda. The Pharmacovigilance Risk Assessment Committee (PRAC) will revise Good Vigilance Practices (GVPs) and other guidance documents. Support will also be provided for the use of artificial intelligence and real-world evidence in regulatory pharmacovigilance activities, reflecting the growing role of data-driven approaches in safety monitoring.

The paediatric drugs

The Paediatric Committee (PDCO) will prioritise making paediatric investigation plans (PIPs) more feasible and ensuring the timely completion of development programmes. Activities will include developing therapeutic area strategies and strengthening cooperation with the Clinical Trials Coordination Group (CTCG). The PDCO will also explore innovative ways to generate and use data for marketing authorisation, including the implementation of the ICH E11 guideline on paediatric extrapolation.

Overall, the EMA work plans for 2026 underline a year shaped by regulatory preparedness, methodological evolution and the progressive integration of digital tools into medicines regulation across the European Union.

EMA-FDA joint principles for the use of AI along medicines lifecycle

0

The European Medicines Agency (EMA) and the US Food and Drug Administration (FDA) have jointly published a document outlining ten principles for the responsible use of artificial intelligence (AI) throughout the medicines lifecycle. The initiative marks a coordinated effort between the two regulators to support the future development of AI guidance across different jurisdictions. The joint principles will provide a common reference framework for the development, validation and oversight of AI systems used in activities related to medicinal products, from early research to post-market phases.

A human-centric system

According to the document, AI systems should be human-centric by design and developed using a risk-based approach. This approach must include appropriate validation processes and risk mitigation measures proportionate to the intended use and potential impact of the system. The development and deployment of AI should comply with applicable legal, ethical, technical, scientific and cybersecurity standards, as well as relevant regulatory requirements, including Good Practices (GxP). A clearly defined context of use is another key requirement. Each AI system should be designed and implemented for a specific purpose, with its scope and limitations clearly documented. Development should involve multidisciplinary expertise across the entire lifecycle of the system, reflecting the complexity of the pharmaceutical regulatory environment. In line with GxP principles, detailed documentation is expected. This includes data source provenance, data processing steps and analytical decisions taken during model development and operation.

Governance and risk-based AI oversight

Robust governance mechanisms must also be in place to ensure the protection of privacy and sensitive information. Data used in AI systems should be fit-for-use, ensuring adequate interpretability, explainability and predictive performance. The agencies stress that performance assessments should follow a risk-based approach and evaluate the AI system as a whole, including human-AI interactions where relevant.The document also calls for the implementation of a risk-based quality management system throughout the technology’s lifecycle. This is aimed at ensuring continuous oversight, reliability and compliance over time. Finally, information generated by AI systems should be presented to users in plain language, ensuring clarity, accessibility and contextual relevance. With these joint principles, EMA and FDA signal a shared commitment to promoting consistent and responsible use of AI in the medicines regulatory space, laying the groundwork for future guidance in this rapidly evolving field.

Newsstand

  • NCF International n.1 - 2026 NCF International n.1 - 2026
  • NCF International n.2 - 2025 NCF International n.2 - 2025
  • NCF International n.1 - 2025 NCF International n.1 - 2025