Home Authors Posts by staff

staff

119 POSTS 0 COMMENTS

The fragilities of the U.S. API supply chain according to the national fragility index

0

A report on the fragility of the U.S. pharmaceutical supply chain published by the API Innovation Center (APIIC) provides new insights into the vulnerabilities affecting the availability of medicines in the United States. Based on the newly developed National Fragility Index (NFI), the analysis found that, in 2023, more than 550 million prescriptions were linked to medicines associated with high or severe supply chain fragility.

The NFI was designed to identify vulnerabilities at the level of individual marketing authorizations across more than 400 products spanning over 130 therapeutic categories. According to Natalie de Graaf, Vice President and General Manager for Data Analytics, AI and ML Solutions for National Security and Health Outcomes, the index provides decision-makers with a practical tool to determine which supply chains require immediate attention and to evaluate whether ongoing mitigation efforts are producing tangible results.

Key Findings from the NFI

According to data released so far by APIIC, 14 medicines have been classified as facing a severe level of supply chain fragility, while approximately 70 additional products require heightened attention. The report also highlights that at least 11 active pharmaceutical ingredients (APIs) included in the U.S. Administration for Strategic Preparedness and Response (ASPR) list of essential medicines have a manufacturer located in the United States. However, none of these manufacturers currently appears to supply U.S.-based pharmaceutical companies.

Furthermore, nearly three-fifths of the medicines analyzed have experienced at least one drug shortage, with an average shortage duration exceeding two years.

The NFI incorporates data from multiple sources and evaluates a broad range of risk factors, including geopolitical considerations, regulatory compliance, financial stability, manufacturing site location and the clinical importance of the products involved. The ultimate goal is to help both public and private investors assess the impact of investments aimed at strengthening domestic manufacturing capacity and reducing supply risks. Cases classified as “severe” require immediate assessment and action, while those rated as “high” call for short-term mitigation measures. APIIC also emphasizes that no pharmaceutical supply chain is entirely risk-free, although vulnerability decreases progressively among products receiving medium or low fragility scores.

Pantoprazole and Metoprolol as Illustrative Cases

Among the examples highlighted in the report is pantoprazole, which received a severe NFI rating. The analysis found that the API was sourced from 36 suppliers across 12 different countries, with no U.S.-based manufacturers involved. At the finished-product level, only 5.6% of the pantoprazole dispensed in the United States was domestically produced.

A similar situation was observed for the beta-blocker metoprolol, for which no U.S. manufacturers were identified. The product has also been affected by two separate shortage events since 2015, each lasting nearly five years.

The National Fragility Index therefore aims to provide a clearer picture of pharmaceutical supply chain vulnerabilities and to support evidence-based investments designed to improve resilience across the U.S. medicines supply network.
When innovation and automation reshape Pharma processes

0

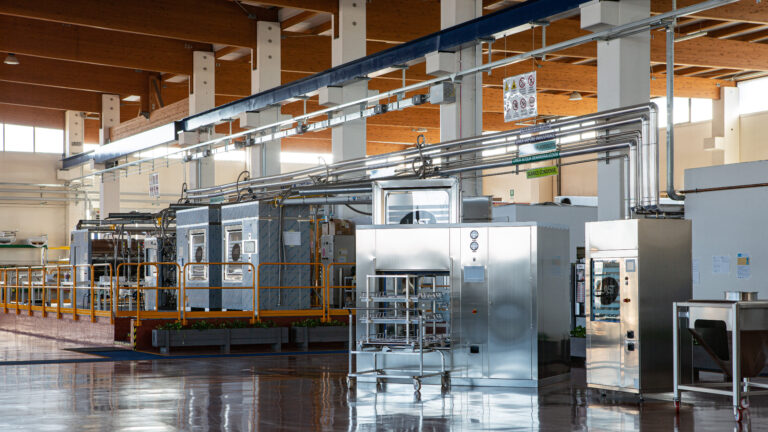
The pharmaceutical industry is undergoing a period of profound transformation, driven by process automation, digital integration and the introduction of increasingly advanced technologies in the management of production stages. In this context, process equipment plays a strategic role in ensuring operational efficiency and maintaining high standards of quality and safety throughout the entire drug production cycle. Among the companies that have contributed to the development of these technologies in recent years is LAST Technology, a firm specialising in the design and manufacture of systems for the pharmaceutical and biopharmaceutical industries and research laboratories. Technological innovation, process digitalisation and sustainability are now some of the cornerstones of the company’s strategy. We spoke about this with Massimo Castellarin, president and CEO of LAST Technology.

R&D is a key element of your development strategy. Can we talk about it?
«We design bespoke solutions, developed to integrate increasingly complex processes and meet specific customer requirements in terms of efficiency, safety and reliability. This translates into the creation of highly customised machines, in which automation and technological integration play an increasingly significant role. A distinctive feature of our business is the use of off-the-shelf components and software, chosen to ensure reliability and design flexibility. This approach allows us to configure equipment in a modular way and facilitates updates over time. The integration between hardware and software also enables the machines to be managed via remote monitoring and control systems, accessible from centralised servers or mobile devices, improving operational efficiency and reducing response times».

What are the main technological innovations you have developed in recent years?
«Among the innovations introduced, we can mention the use of acetone in washing processes, a particularly effective innovation for removing stubborn residues associated with pharmaceutical active ingredients. Then there are the “combo” solutions that integrate washing and sterilisation into a single process, particularly useful for elastomers and solid products, with advantages in terms of operational simplification and time savings. We have also introduced combined sterilisation systems using superheated water and steam, designed to treat both solids and liquids in rigid or flexible containers. For heat-sensitive components, low-temperature sterilisation technologies are available, based on the use of ethylene oxide or hydrogen peroxide. Finally, we have developed hot-air drying machines with an ATEX system, designed for the safe processing of potentially explosive solid products».

How do your solutions fit into the “automated factory” model?
«One of the most significant aspects concerns the integration of automation systems into handling and materials management processes. Today, it is possible to automate various operational phases through the use of LGV shuttles, anthropomorphic robots for pick and place operations, and automatic feed-in and feed-out systems, which reduce the need for manual intervention and improve the continuity of the production process. At the same time, we are working on integrating the equipment into digital plant supervision systems. Our machines can be connected to SCADA platforms for process monitoring and integrated into corporate MES systems, becoming part of a fully interconnected production ecosystem. To support this model, we have also developed proprietary software, based on standard market communication protocols, which allows all LAST Technology process machines to be incorporated into the corporate IT domain».

Does innovation also extend to the organisation of business processes, is that correct?
«In this regard, we have improved order management, making the entire product lifecycle more efficient. In a complex sector such as pharmaceutical equipment, the ability to coordinate information, technical documentation and operational activities is a strategic factor. This is why we have adopted a PLM (Product Lifecycle Management) platform that enables the integration of data, applications and operational methodologies throughout all project phases, from the concept design stage right through to plant commissioning and qualification (C&Q). This system allows the various company departments involved in order management to be interconnected, improving collaboration and information sharing. The introduction of PLM has reduced order management times by 20-30%, whilst also increasing the level of control over processes and technical documentation. To support operational activities, we have also introduced mobile IT workstations in the assembly and testing areas, enabling internal staff and visiting clients to quickly access technical documentation relating to individual orders. This approach helps to make processes more transparent, efficient and integrated with the quality management system».

Sustainability: a real goal

«Sustainability is an integral part of our corporate strategy and has been progressively incorporated into decision-making processes and development models – Castellarin explains– We draw inspiration from the United Nations Sustainable Development Goals (SDGs) with the aim of creating shared value for all stakeholders, integrating economic growth, social responsibility and attention to environmental impact. In recent years, we have launched several concrete initiatives in this direction. This commitment has earned the company the EcoVadis Silver Medal, one of the leading international systems for assessing sustainability performance. We have taken steps to improve the plant’s energy efficiency and energy supply. With this in mind, we have installed a 515 kWh photovoltaic system which, in addition to enabling self-sufficiency, has significantly reduced emissions. Our goal is to achieve carbon neutrality by 2030. Looking to the future, we will continue to invest in the development of increasingly efficient and sustainable technologies, whilst keeping a close eye on industry developments, where automation and artificial intelligence will play an ever-greater role in improving the efficiency and quality of production processes».

Cancer medicines: faster approvals, but access remains uneven across Europe

0

A new report from the European Federation of Pharmaceutical Industries and Associations (EFPIA), prepared by the Swedish Institute for Health Economics (IHE), highlights continued disparities in access to innovative cancer treatments across Europe. While regulatory approvals have accelerated significantly, many patients still face delays before new medicines become available in clinical practice. According to the report, cancer could become the leading cause of death in the European Union by 2035. However, several countries have already recorded a decline in cancer mortality rates over the past decades.

Access gap persists despite regulatory progress

The report suggests that centralized approval by the European Medicines Agency (EMA) is no longer the main bottleneck. Instead, delays often occur after authorization, when medicines must be incorporated into national, regional, or local reimbursement systems, formularies, and clinical guidelines. Countries such as Italy, Spain, and Sweden are cited as examples where additional regional approval processes can slow patient access to newly approved therapies.

Between 1995 and 2024, EMA approved 194 oncology medicines, alongside more than 300 new indications for existing products. Yet access remains highly uneven. Western European countries generally provide broader and faster reimbursement than countries in Central and Eastern Europe. In some smaller markets, including Malta, Estonia, Slovakia, and Romania, only around 20% of cancer medicines approved between 2019 and 2022 had secured reimbursement. In Latvia and Lithuania, reimbursement procedures can take up to 900 days.

Structural barriers still limit uptake

The report identifies several factors that continue to hinder the adoption of innovative oncology medicines across Europe. These include differing pricing and reimbursement systems, variations in health technology assessment (HTA) requirements, insufficient reimbursement for companion diagnostics, and limited molecular diagnostic infrastructure in some countries.

Despite these challenges, the gap in the uptake of new cancer medicines has narrowed in recent years. The difference between the highest- and lowest-performing countries fell from 5.3-fold in 2018 to 3.3-fold in 2023. Austria, France, and Switzerland are among the fastest adopters of new oncology therapies in clinical practice.

Recommendations for a more equitable system

To improve access, the report calls for greater use of managed entry agreements (MEAs), including outcome-based arrangements, and more flexible HTA pathways that prioritize medicines with demonstrated clinical value. Additional measures include wider use of biomarker-driven patient stratification, optimization of treatment sequencing, and greater adoption of generic and biosimilar medicines.

The authors also highlight the need for stronger real-world evidence collection through disease registries to support healthcare decision-making. Looking ahead, Europe’s evolving pharmaceutical legislation and the implementation of the EU HTA Regulation could help accelerate reimbursement decisions and encourage broader availability of innovative cancer medicines across all Member States. However, questions remain around national acceptance of joint clinical assessments and Europe’s competitiveness as a destination for oncology clinical research.

EMA identifies ten priority actions to advance women’s health

0

A new regulatory roadmap aimed at improving how women’s health is addressed in medicines development and regulation is expected to be published by the end of 2026. The initiative was announced by the European Medicines Agency (EMA) following a multi-stakeholder workshop held in Amsterdam on 28-29 September, which brought together more than 100 participants from across the healthcare ecosystem.

“Women are still missing: from research and data to prevention, diagnosis and treatment,” said EMA Executive Director Emer Cooke. “But the timing for improving support for women’s health has never been better. Over the last two days, we have seen what can be achieved when diverse voices unite behind a common cause. Now is the time to act. EMA will build on this dialogue to shape a roadmap for medicines addressing women’s health that will help drive progress in Europe and beyond.”

The proposed actions

The Amsterdam workshop identified ten priority actions across areas where EMA can lead, collaborate or influence the development of products for women’s health. While details will be outlined in the forthcoming roadmap, the broad priorities have already been defined. These include stronger engagement with patients and stakeholders to identify priorities, unmet needs and existing gaps, alongside enhanced support for research activities, including funding opportunities. Participants also highlighted the need to improve women’s recruitment into clinical trials, including pregnant and breastfeeding women, while ensuring the collection and reporting of high-quality sex-disaggregated data.

Another priority is to generate evidence throughout the entire lifecycle of medicines by ensuring the inclusion of women of all ages and life stages in research. Greater use of real-world data, supported by sustainable infrastructure, was also identified as a key enabler. At the regulatory level, the roadmap is expected to introduce training initiatives on sex- and gender-sensitive assessments across the European medicines regulatory network. It will also focus on improving communication of benefits, risks and uncertainties related to medicine use in women.

Finally, the roadmap is set to include measurable outcomes, key performance indicators (KPIs) and structured international collaboration to track progress and promote a more gender-responsive approach to medicines development and regulation.

Image generated with AI

The Advocate General’s opinion on the EPR awaiting the Court’s final decision

0

The European associations representing the pharmaceutical and consumer healthcare sectors have welcomed the opinion issued on 3 September by the Advocate General in Case C‑193/25 before the Court of Justice of the European Union. The organizations, including EFPIA, Medicines for Europe and AESGP, view the opinion as an important step in addressing concerns over the allocation of costs under the revised Urban Wastewater Treatment Directive (UWWTD).

The case stems from a challenge brought by Poland, which asked the Court to assess whether the directive’s Extended Producer Responsibility (EPR) provisions are consistent with the polluter-pays principle and the principle of equal treatment. Under the current rules, manufacturers of pharmaceuticals and cosmetics are required to finance at least 80% of the costs associated with quaternary wastewater treatment.

Concerns over the scientific basis

In his opinion, the Advocate General recommended that the Court annul Article 9(1)(a) and Annex III of the UWWTD, the provisions establishing the financing obligations for pharmaceutical and cosmetic companies. According to the opinion, the EU institutions did not sufficiently demonstrate that the allocation of responsibility was supported by a robust assessment of the available scientific evidence. Particular attention was given to the methodology used to calculate the toxic load attributed to different sectors, with the Advocate General identifying what were described as significant shortcomings in the underlying approach. Industry associations argue that these findings reinforce long-standing concerns regarding the evidence base used to justify the financial burden placed on the pharmaceutical and cosmetics industries.

Call for a temporary suspension

Following the publication of the opinion, the associations called on the European Commission to temporarily suspend implementation of the EPR scheme until the Court of Justice delivers its final ruling. According to the industry groups, pausing implementation would provide legal certainty while the judicial process is ongoing and would avoid potentially irreversible consequences should the Court ultimately decide to annul the contested provisions. The final judgment of the Court is now expected to determine the future of the EPR framework under the UWWTD and could have broader implications for how environmental responsibilities are allocated across industrial sectors within the European Union.

Global regulators call for broader adoption of alternatives to animal testing

0

The International Coalition of Medicines Regulatory Authorities (ICMRA) has issued a joint statement urging stakeholders across the pharmaceutical value chain to accelerate the development, validation and regulatory implementation of alternative methods to animal testing wherever scientifically appropriate. The initiative, published on 1 October and endorsed by the European Medicines Agency (EMA), reflects a growing international consensus that advances in science can support a progressive reduction in the use of animals while maintaining high standards of product quality, safety and efficacy.

The statement is addressed to regulators, pharmaceutical developers and manufacturers, as well as national control laboratories. Its objective is to provide a shared regulatory perspective on the adoption of so-called New Approach Methodologies (NAMs) and to encourage their consistent implementation across global jurisdictions. The document was developed under the leadership of EMA, the UK’s Medicines and Healthcare products Regulatory Agency (MHRA), the Irish Health Products Regulatory Authority (HPRA) and Swissmedic.

A lifecycle-wide approach

According to ICMRA, scientific progress over recent years has created new opportunities to reduce reliance on animal testing throughout the entire medicine lifecycle. The coalition highlights the potential application of alternative approaches from early-stage candidate screening and preclinical development through to clinical studies, marketing authorisation and post-marketing activities.

The statement notes that international regulatory frameworks already provide a foundation for this transition. Existing International Council for Harmonisation (ICH) guidelines support the application of the 3Rs principles (Replacement, Reduction and Refinement) and outline key considerations for the development of non-animal methods. Further updates to existing guidelines, along with the creation of new ones, could help clarify regulatory expectations and facilitate broader adoption of innovative testing approaches.

ICMRA points out that many regulatory authorities already recognise that there is often no scientific justification for conducting animal studies for generic medicines and biosimilars. The same principle may apply when a drug’s pharmacological activity cannot be meaningfully reproduced in available animal species.

The coalition suggests that, where supported by robust scientific evidence, similar approaches could be extended to other medicinal products, including those that are pharmacologically active in animals.

A cautious transition

While advocating for greater use of alternative methods, regulators stress that the transition must be guided by scientific validity and regulatory acceptability. The statement emphasises that replacing animal studies requires a careful and evidence-based approach to ensure continued protection of patient safety.

According to ICMRA, the complete elimination of animal testing for areas such as general toxicity, reproductive toxicity, carcinogenicity, pharmacokinetics and pharmacodynamics remains a long-term objective rather than an immediate goal.

The coalition expects one of the most significant near-term impacts of alternative methods to come from their wider use in routine batch release testing. In this area, in vivo testing could increasingly be restricted to situations where strong scientific evidence supports its necessity and where no equivalent or superior alternative methods are available.

Supporting global implementation

ICMRA acknowledges that different countries and regulatory authorities are likely to move at different speeds in adopting non-animal methods. As a result, the organisation is calling on pharmaceutical companies, developers and manufacturers to actively support the transition and collaborate with regulators to accelerate implementation.

Possible incentives highlighted in the statement include dedicated priority meetings and fee waivers for scientific advice related to the qualification and regulatory acceptance of new methodologies.

The joint declaration represents another step in the global effort to modernise drug development and regulatory assessment, balancing scientific innovation, animal welfare considerations and the need to maintain confidence in the safety and quality of medicines.

ePI: EMA publishes guidance for submission in centralised procedures

0

Preparatory activities are continuing in anticipation of the submission and publication of electronic product information (ePI), including electronic package leaflets and other product information documents, as part of the broader implementation of the new European pharmaceutical legislation.

The European Medicines Agency (EMA) has published guidance on the procedures to be followed for ePI relating to centrally authorised products. The guidance applies exclusively to applications submitted after the planned implementation “go-live” date (see the implementation roadmap). EMA emphasizes that this new guidance does not replace any of the regulatory documents currently in force.

Key guidance on ePI submission and publication

Electronic Product Information (ePI) encompasses the Summary of Product Characteristics (SmPC), labelling, and the package leaflet. For centrally authorised medicinal products, submission of these documents in electronic format will remain voluntary until the new Directive on medicinal products for human use becomes fully applicable. During this transitional phase, ePI may be submitted in English, while submission in other EU languages, as well as Icelandic and Norwegian, remains optional. This situation may change in the future, with mandatory submission of ePI in all relevant languages. Once published, electronic product information must be kept aligned with the latest approved version under the responsibility of the applicant.

EMA’s guidance provides instructions on when ePI should be submitted through the Product Lifecycle Management (PLM) Portal, in parallel with the various regulatory procedures affecting a medicinal product throughout its lifecycle. It also highlights that electronic product information, including any subsequent versions, is always linked exclusively to the product’s marketing authorisation number. The connection between the ePI and the medicinal product authorisation must be established through the PLM Portal before ePI submission. The guidance then describes the entire process through to publication of the ePI. Specific recommendations are also provided for small and medium-sized enterprises (SMEs) submitting ePI as applicants.

Creation and update of electronic product information

A dedicated section is devoted to the initial creation of electronic product information, both within the context of a new marketing authorisation application and for products that have already been authorised. The guidance also explains the process for updating ePI following its initial publication and outlines how different post-authorisation procedures may affect those updates. Finally, the document provides guidance on the management of ePI in cases involving suspension, transfer, withdrawal, or refusal of a marketing authorisation, as well as changes to a medicinal product’s brand name.

EMA launches pilot project for voluntary submission of NAM data

0

Among the first developments following the summer break, the European Medicines Agency (EMA) has launched a new pilot project on the voluntary submission of data (the Voluntary Data Submission, VDS, pilot) generated using New Approach Methodologies (NAMs). These include in vitro, in silico and in chemico methods, as well as refined ex vivo and in vivo approaches designed to progressively reduce reliance on traditional animal testing.

According to EMA, the call for NAM data submissions is expected to remain open until the third quarter of 2027, with the final outcomes of the pilot scheduled for publication by the end of next year.

A pilot to support data sharing

The “safe harbour” mechanism for voluntary data sharing was originally introduced in 2012 through guidance on the regulatory acceptability of testing approaches based on the 3Rs principles (replacement, reduction and refinement). However, industry uptake has so far been limited. Companies were given the opportunity to voluntarily submit data from NAM studies alongside data generated through conventional approaches as part of marketing authorisation applications, with the aim of helping regulatory authorities gain practical experience with these emerging methods.

The recently published EMA document highlights that experience with NAMs remains very limited among European regulators. Fundamental questions regarding the feasibility, incentives, data management requirements and regulatory added value of these novel preclinical assessment methodologies also remain largely unexplored.

The new VDS pilot significantly redesigns the approach to voluntary NAM data sharing by decoupling it from marketing authorisation procedures. Through this initiative, EMA aims to gain practical experience, identify scientific and operational challenges, and assess the feasibility of developing a more structured VDS framework in the future. Among the Agency’s key objectives are improving understanding and confidence in the evaluation of NAM data and providing a platform for discussion of NAM-related proposals, including their Contexts of Use (CoU) and Regulatory Applicability Areas (RAA).

EMA intends to provide participating organisations with non-binding scientific feedback on the regulatory relevance and fitness for purpose of the submitted methods, together with recommendations on possible next steps for further discussion through mechanisms such as the Innovation Task Force (ITF), Scientific Advice (SA) or Qualification Advice (QA) procedures.

The initiative is also expected to contribute to a harmonised interpretation of NAM data and Weight of Evidence (WoE) approaches across the EU regulatory network.

How to participate in the pilot

The pilot project is open to pharmaceutical companies, contract research organisations (CROs), method developers, non-governmental organisations, academic laboratories and consortia. Interested parties may submit an expression of interest to EMA using the dedicated application form available on the Agency’s website.

Four priority Regulatory Applicability Areas (RAAs) have been identified for participation in the pilot:

  • Risk assessment for biological medicinal products lacking a pharmacologically relevant animal species;
  • Developmental and reproductive toxicity (ICH S5(R3));
  • Safety pharmacology (ICH S7A);
  • Hepatotoxicity and drug-induced liver injury (DILI).

Following its internal assessment, EMA may redirect applicants to other, more appropriate procedures, such as ITF, SA or QA. Selected participants will be invited to submit a more comprehensive briefing package and will be provided with an EMA account and access to EudraLink.

Data will be shared through EudraLink and securely stored within EMA’s SharePoint system, accessible only to the Operational Expert Group (OEG) and relevant Agency staff. The briefing package should be aligned, in terms of format and level of detail, with Module 2.4 of the electronic Common Technical Document (eCTD). It should include a description of the RAA, the endpoints of interest, internal validation criteria and the intended regulatory application of the proposed methodology.

AI-generated image by the Editorial Team

EMA maps pharma’s sustainability pathway

0

The event, coordinated by the Quality Innovation Group (QIG) and its Listen and Learn Focus Group (LLFG), brought together industry representatives, academics and regulators, including the European Commission, EDQM, FDA, PMDA and Swissmedic. The main conclusion was that sustainability can be integrated into pharmaceutical development and manufacturing through a gradual, science-based approach that preserves product quality, safety and efficacy.

Among the case studies presented, a consortium led by Imperial College London showed how digital tools combining techno-economic modelling and Life Cycle Assessment (LCA) can support evidence-based sustainability decisions. Researchers highlighted applications ranging from lentiviral vector production to monoclonal antibody manufacturing, while also pointing to the need for shared, non-confidential datasets to improve sustainability benchmarking.

Packaging and energy under the spotlight

Packaging emerged as another major focus area. Discussions covered the potential adoption of bio-based drop-in plastics, chemically identical to fossil-derived polymers, which could simplify regulatory implementation if supported by appropriate risk assessments. Participants also examined pathways to replace PVC blister packs, with industry groups proposing harmonised methods based on water vapour transmission rate (MVTR) to reduce the burden of product-specific stability studies.

Energy efficiency in manufacturing was another recurring theme. Data presented by GSK indicated that HVAC systems can account for up to 60% of a pharmaceutical site’s energy consumption. Adaptive HVAC technologies capable of adjusting airflow in real time could reduce energy use by more than 30% in Grade C and D environments, provided qualification and monitoring requirements can be adequately addressed.

Green chemistry and alternative methods

The Horizon Europe IMPACTIVE project showcased mechanochemistry as a viable alternative to solvent-intensive synthesis processes. One case study reported an 85% reduction in CO₂ emissions and environmental toxicity, together with lower production costs, through the use of twin-screw extrusion for nitrofurantoin manufacturing.

Additional discussions focused on predictive models for solvent recovery and recombinant alternatives to traditional bacterial endotoxin testing. Industry representatives highlighted recombinant Factor C (rFC) and recombinant Cascade Reagent (rCR) methods as more sustainable alternatives to tests relying on horseshoe crab-derived reagents.

Looking ahead, workshop participants agreed that sustainability should not become a standalone fourth regulatory pillar alongside quality, safety and efficacy. Instead, regulators and industry will continue working on targeted guidance, scientific alignment and international collaboration to facilitate the adoption of sustainable technologies across the pharmaceutical sector.

IMA Pharma and IMA MedTech at CPHI Milan 2026

0

CPHI Worldwide brings together the entire pharmaceutical supply chain and stands as one of the most important events in the industry’s annual agenda. The Pharmaceutical and MedTech Divisions of IMA Group, together with their specialists, are pleased to guide you inside Pharma, introducing the All-In-One supplier specialized in designing and manufacturing innovative machines and complete lines for the processing and packaging of pharmaceutical, nutraceutical and medical products. With a strong focus on innovation, IMA also delivers projects dedicated to digitalisation, sustainability, and customer service solutions, supporting the entire pharma ecosystem towards greater efficiency and responsibility. Here is a preview of what you will find on display.

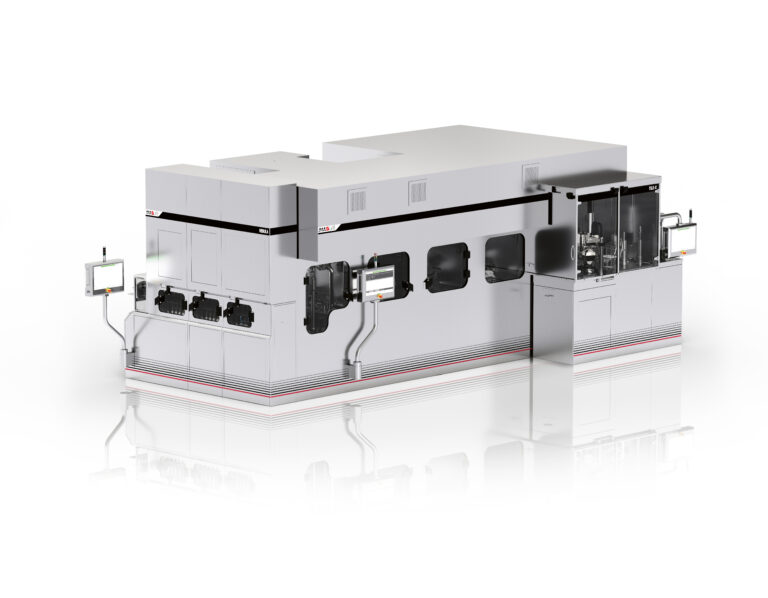
Following its world premiere at Interpack 2026, IMA Life will present the TILE-X line at CPHI Milan, where it will make its Italian debut. TILE-X represents a groundbreaking innovation in the fill-finish process, designed to handle Ready-To-Use components through magnetic levitation technology. Suitable for high-value pharmaceuticals such as biopharmaceuticals and ATMPs with a focus on Cell & Gene therapies, TILE-X extends the concept of modularity, unprecedented in the aseptic fill-finish process. Based on magnetic levitation technologies, TILE-X enables frictionless component handling with zero risk of particulate generation.

At the IsoTech Lab – a corner devoted to the Isolator Research & Development Laboratory – IMA Life experts in aseptic processing will present the new IMA VPHP BioIndicator. Developed to complement Microbiological Support Services, this new generation of biological indicators has been specifically designed for VPHP bio-decontamination cycles used in isolators. IMA VPHP BioIndicator supports Cycle Development, Performance Qualification (PQ), re-qualification, and ongoing contamination control strategies, ensuring robust and defendable 6-log sporicidal performance. These biological indicators are based on Geobacillus stearothermophilus spores (ATCC 12980) and packaged in VPHP-permeable packaging.

 VPHP BioIndicator, D-value studies

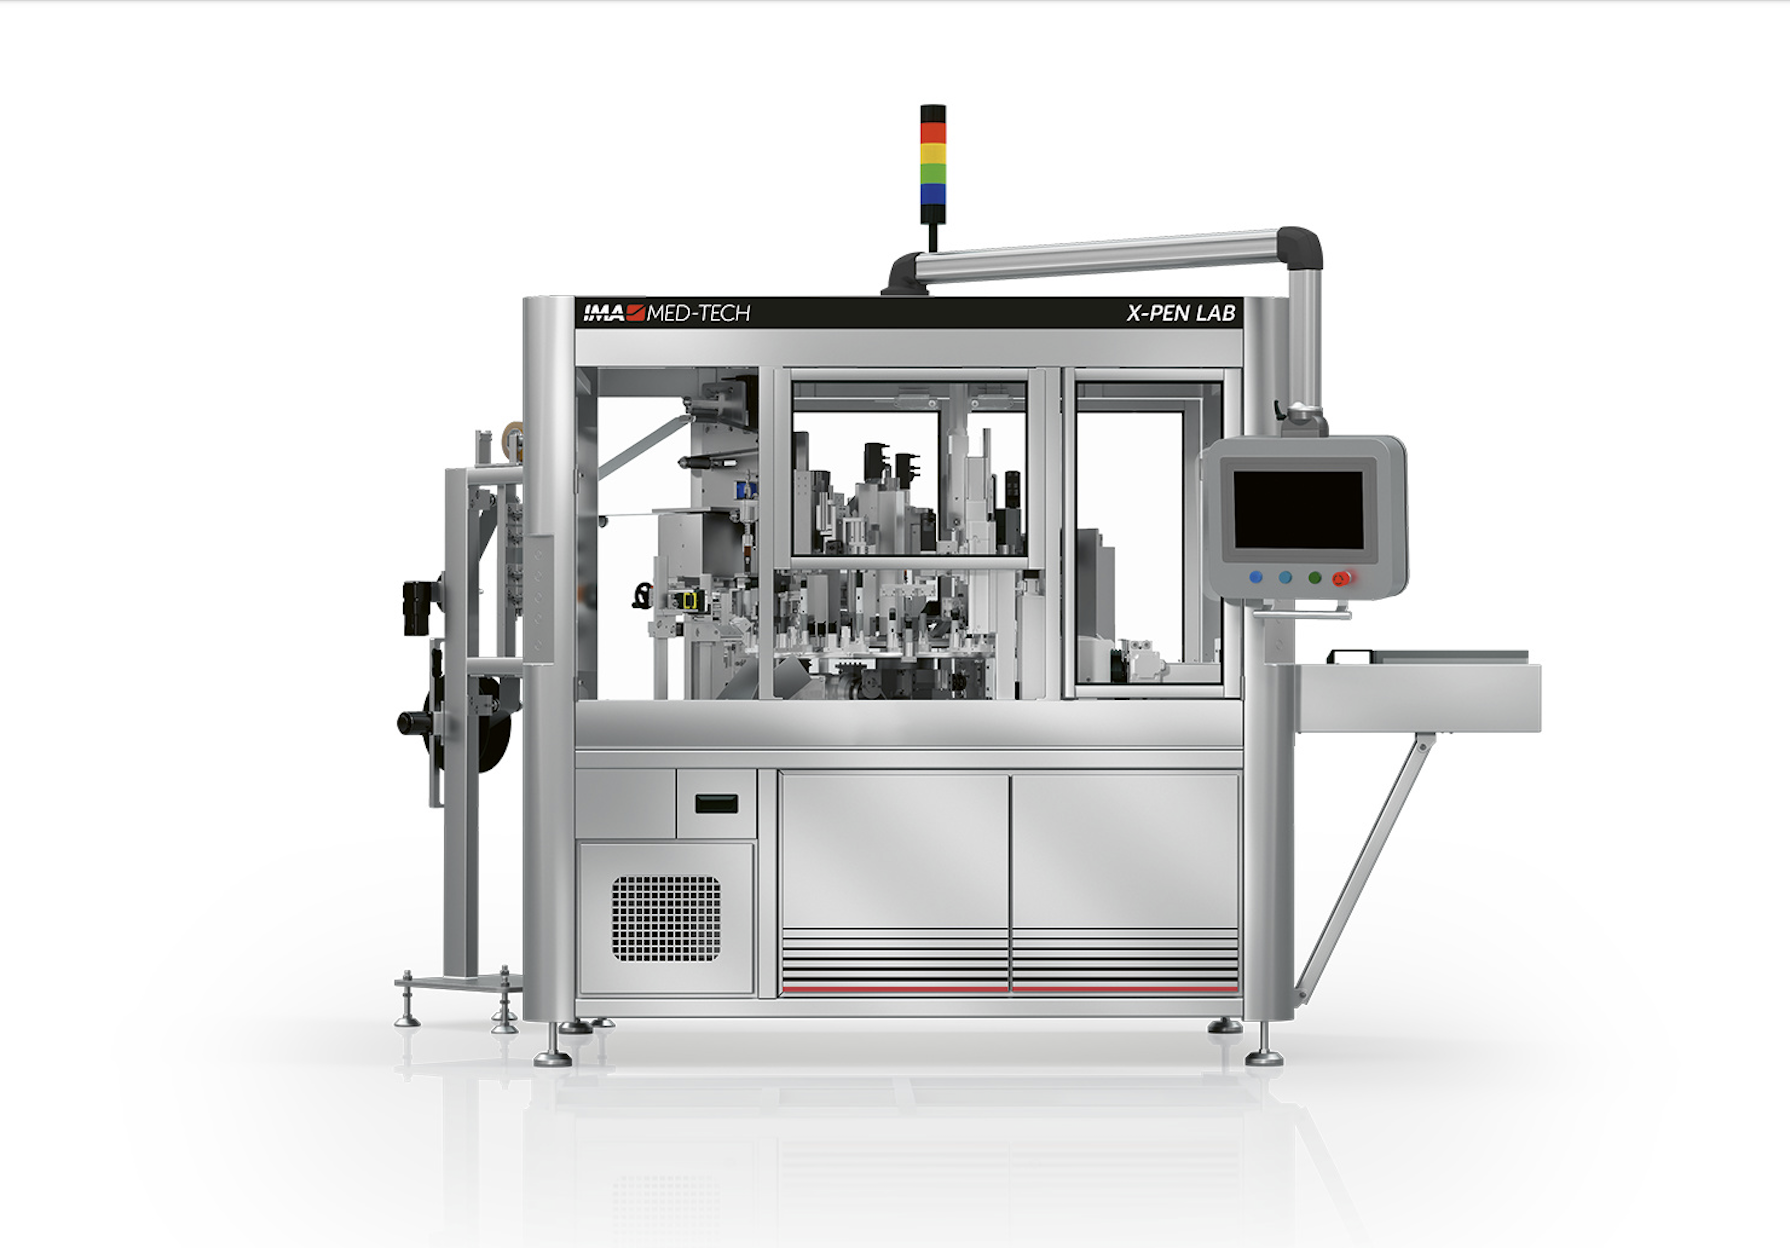
On show also the X‑PEN LAB, a precise semi‑automatic solution for clinical trials and small‑scale production. It ensures accurate press‑in with monitored force‑distance measurement and supports multi‑variant manufacturing with manual loading and offloading. Ideal for early‑stage development, X‑PEN LAB belongs to the X‑PEN Series: IMA MED‑TECH’s standard platform for pen and autoinjector final assembly. The X‑PEN platform, based on a Linear Magnetic Transport System (LMTS), supports multi‑variant manufacturing with production speeds ranging from 40 to 160 ppm. Integrated options include vision inspection, no‑glass‑to‑glass PFS handling, force‑distance controlled assembly, priming and labelling.

X-PEN LAB 

For further information:

Newsstand

  • NCF International n.2 - 2026 NCF International n.2 - 2026
  • NCF International n.1 - 2026 NCF International n.1 - 2026
  • NCF International n.2 - 2025 NCF International n.2 - 2025